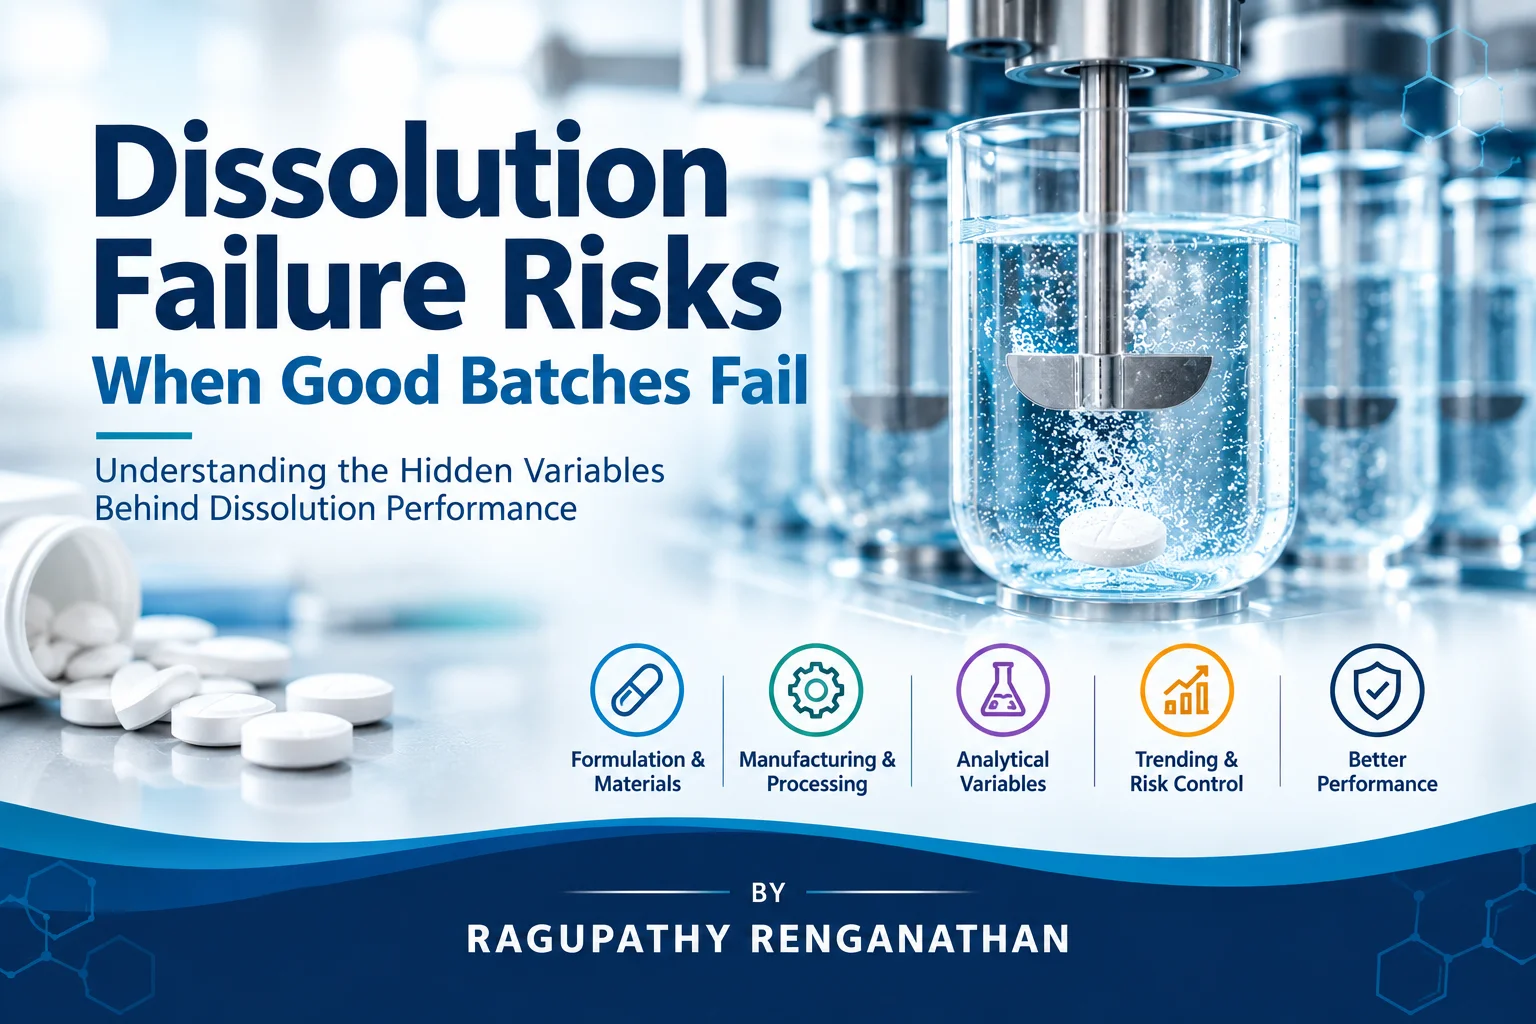
Understanding the Hidden Variables Behind Dissolution Performance
In pharmaceutical manufacturing, a batch can meet assay requirements, pass content uniformity, satisfy physical specifications, and appear completely acceptable on the production floor—yet still fail dissolution. This is one of the most important realities of oral solid dosage manufacturing: a product can look like a “good batch” from several conventional quality perspectives while its drug-release behavior reveals a deeper formulation, material, process, or analytical problem.
Dissolution testing is not simply another laboratory test performed at the end of manufacturing. It is a performance-oriented quality attribute that provides information about how the drug substance is released from the finished dosage form. FDA describes the objective of in-vitro dissolution testing as evaluating variables affecting the rate and extent of drug release and, where appropriately designed, helping ensure consistent product quality and performance.
This is why dissolution failures deserve a fundamentally different mindset. A failed result should not immediately be interpreted as a single compression problem, a laboratory error, or a manufacturing deviation. It can be the visible consequence of subtle changes occurring much earlier in the product lifecycle—from API particle-size distribution and polymorphic form to excipient functionality, granulation moisture, milling, lubrication, compression force, coating behavior, equipment hydrodynamics, dissolution-media preparation, sampling, filtration, or analytical measurement.
The complexity becomes even greater when a batch is only marginally below its dissolution specification. A result that passes comfortably in one batch but approaches the lower acceptance boundary in another may indicate a process that remains technically capable of producing acceptable product but lacks sufficient robustness. For pharmaceutical manufacturers, that distinction matters. The objective should not simply be to manufacture batches that pass; it should be to understand and control the variables that consistently produce the intended dissolution profile.
When a Compliant Batch Becomes a Dissolution Risk
Dissolution performance begins with the drug substance itself. The API's aqueous solubility, particle-size distribution, crystal form, surface characteristics, wettability, and other material attributes can influence how rapidly it becomes available for dissolution. Two API lots can meet the same chemical specifications while exhibiting physical differences that influence finished-product performance. FDA inspection guidance has historically highlighted the importance of raw-material physical characteristics, including particle-size profiles, and has documented cases where changes in granulation particle size contributed to dissolution failure.
Particle size is particularly important because it can influence surface area, dissolution kinetics, blend behavior, granulation characteristics, and ultimately tablet microstructure. A smaller API particle may present greater surface area to the dissolution medium, while larger particles may dissolve more slowly under otherwise comparable conditions. However, particle size should never be considered in isolation. Changes in particle morphology, agglomeration, surface energy, crystal habit, or polymorphic form can also alter wetting and dissolution behavior.