Introduction
An Abbreviated New Drug Application (ANDA) is the regulatory pathway used in the United States for qualifying generic drug products. Unlike an NDA, an ANDA generally relies on FDA’s previous finding of safety and effectiveness for the reference listed drug (RLD), while the applicant must provide the information required to demonstrate, among other things, pharmaceutical equivalence, bioequivalence, and the identity, strength, quality, and purity of the proposed product.
Preparing an ANDA is therefore much more than assembling documents. The submission needs to be scientifically justified, internally consistent, appropriately formatted, and complete across areas such as CMC, bioequivalence, labeling, microbiology, administrative information, facilities, and regulatory requirements.
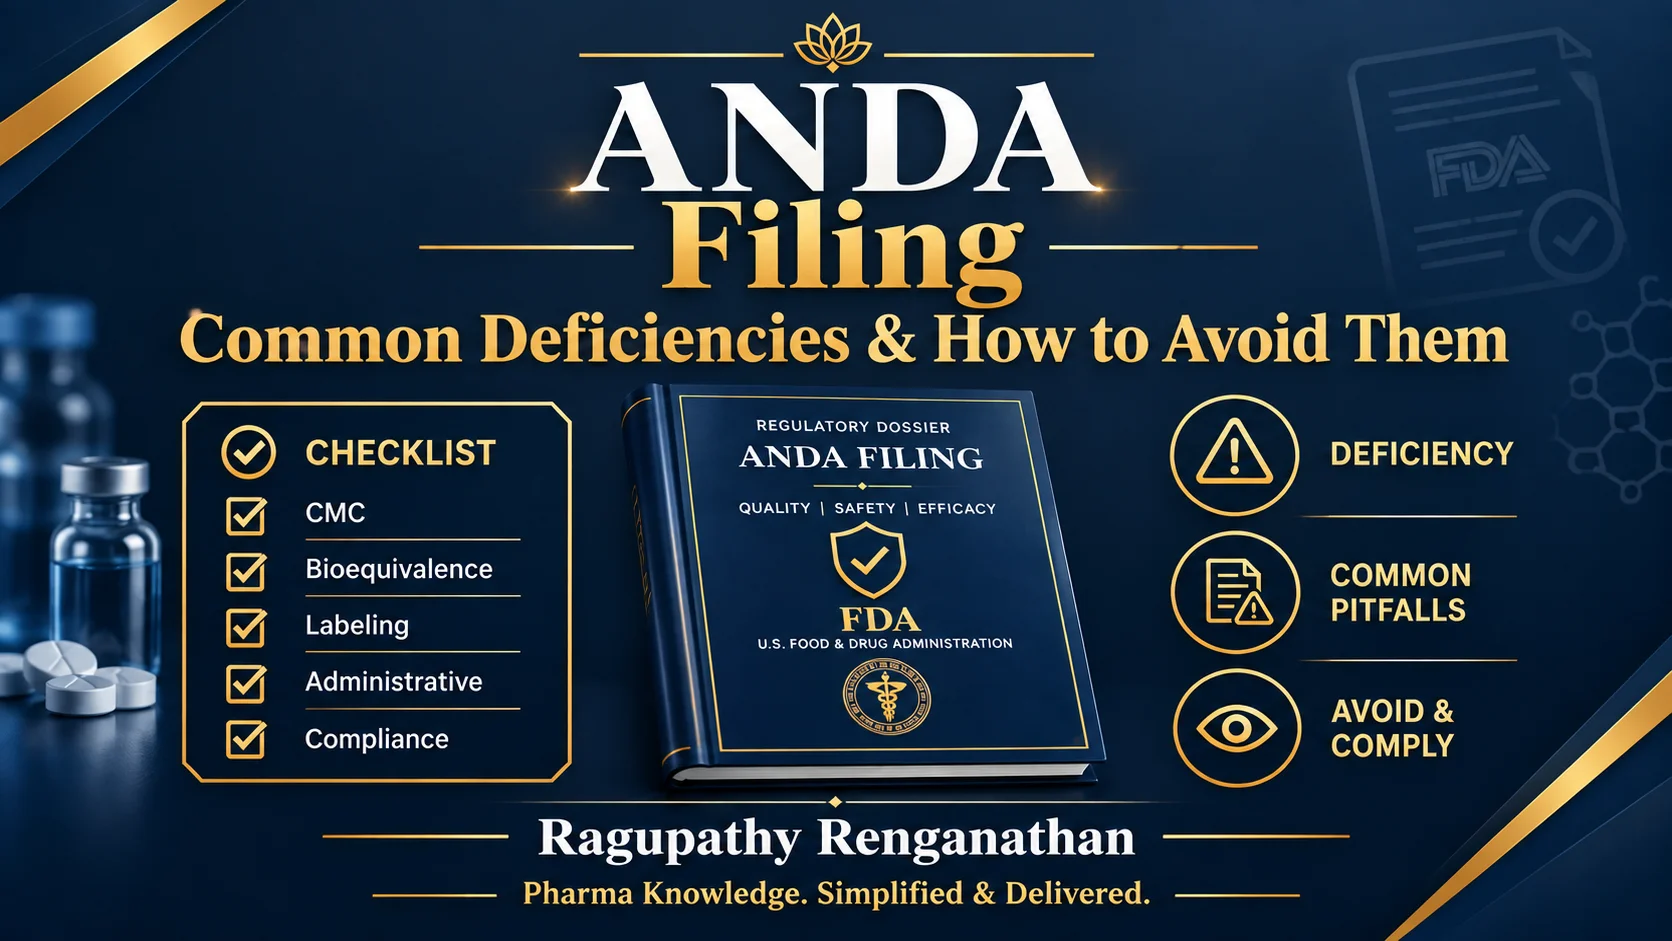
FDA specifically identifies recurring deficiencies that can delay approval, and its Good ANDA Submission Practices guidance is intended to help applicants reduce avoidable deficiencies and minimize review cycles.
A particularly important distinction is between deficiencies identified during filing review and deficiencies identified during substantive review. An application that is not sufficiently complete may be refused to receive (RTR), while deficiencies identified during substantive review may ultimately be communicated through a Complete Response Letter (CRL).
1. Incomplete or Inconsistent Administrative Information
One of the most avoidable problems in an ANDA is inconsistency in basic application information.
Applicant details, manufacturing sites, drug product information, strengths, dosage forms, correspondence information and other administrative elements need to be accurate and consistent throughout the submission.
How to avoid it
Establish an administrative checklist before submission. Cross-check the information across the eCTD, forms, cover letters, facility information, labeling and supporting documentation. A final cross-functional review can identify inconsistencies before FDA review begins.
2. Poorly Organized ANDA Submission
An ANDA is submitted using the Common Technical Document (CTD) structure. Poor organization can make important information difficult to locate and may contribute to filing deficiencies.
How to avoid it
Use the applicable FDA content-and-format recommendations and maintain a clear document structure. Ensure that documents are placed in the appropriate CTD sections and that references between sections are accurate. FDA's ANDA content-and-format guidance specifically describes the information expected in each CTD section.
3. Inadequate CMC Information
Chemistry, Manufacturing and Controls (CMC) information is one of the central components of an ANDA.
The submission needs to provide adequate information concerning the drug substance, drug product, manufacturing process, controls, specifications, analytical procedures and other applicable quality attributes.
How to avoid it
Perform a complete CMC gap assessment before submission. Confirm that manufacturing processes are sufficiently described and that specifications and analytical controls are scientifically justified and appropriately supported by data.
4. Weak Pharmaceutical Development Rationale
A formulation should not simply be presented as a collection of ingredients and manufacturing steps.
FDA expects applicants to provide appropriate information supporting the development and control of the proposed product. The scientific rationale should connect formulation composition, manufacturing variables and product quality.
How to avoid it
Clearly document the development rationale, critical quality considerations, formulation selection and relevant development studies. Ensure that the final control strategy is consistent with the development knowledge.
5. Inadequate Impurity Assessment
Impurities can become an important regulatory issue when their identity, levels, specifications or justification are not adequately addressed.
FDA has specifically issued guidance concerning ANDA submissions that may be refused to receive because of inadequate justification of impurity limits.
How to avoid it
Identify relevant process-related and degradation impurities, establish appropriate controls, provide supporting analytical data and scientifically justify impurity specifications where required.
6. Inadequate Analytical Method Information
Analytical procedures are fundamental to demonstrating product quality.
Problems can occur when methods are inadequately described, insufficiently validated, poorly justified or inconsistently reported across different sections of the application.
How to avoid it
Ensure that analytical methods are appropriately developed, validated and documented according to their intended use. Cross-check method descriptions, specifications and validation information throughout the ANDA.
7. Inconsistent Specifications
Specifications appearing in different parts of the submission should not contradict one another.
Differences in acceptance criteria, test methods, terminology or units can raise questions during regulatory review.
How to avoid it
Create a controlled specification matrix covering raw materials, intermediates, drug substance, drug product and relevant packaging or microbiological controls. Perform a final consistency review before submission.
8. Inadequate Stability Data
Stability information supports the proposed shelf life and storage conditions of the drug product.
Insufficient data, inconsistent stability protocols, unexplained trends or inappropriate interpretation can create regulatory questions.